
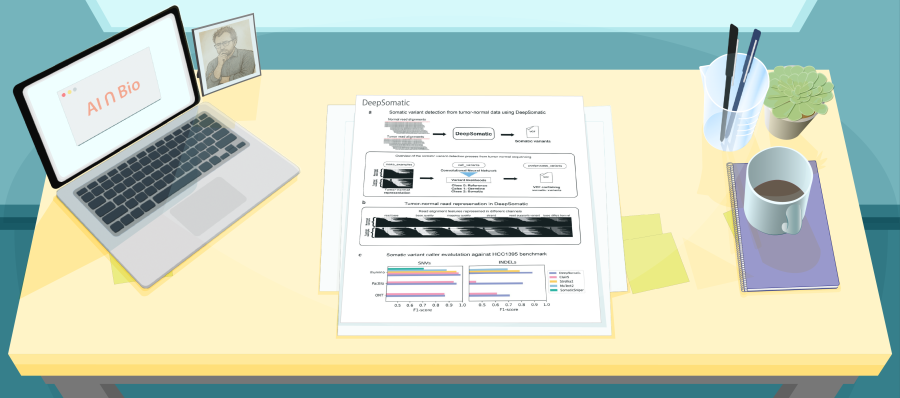
DeepSomatic, published this month in Nature Biotechnology, represents a milestone for cancer genomics: a deep-learning method that detects somatic small variants across both short- and long-read sequencing data. Built on Google’s DeepVariant framework, it bridges Illumina, PacBio HiFi, and Oxford Nanopore datasets and introduces CASTLE, a new multi-platform benchmark of six tumor–normal cell lines made openly available to the community. For anyone working in precision oncology, the technical ambition here is remarkable: one model spanning technologies, sample types, and variant classes.
Scientific Insight
DeepSomatic converts paired tumor–normal reads into tensor “images” that feed a convolutional neural network capable of distinguishing somatic, germline, and reference variants. The model outperformed leading tools such as Strelka2 and ClairS across variant types and variant allele frequencies, and it maintained accuracy across multiple sequencing chemistries. Beyond its raw performance, the CASTLE dataset fills a major gap in the field: creating a real benchmark for long-read somatic variant detection where none previously existed.
Scientific Rigor Note
Like many GenAI systems, DeepSomatic may fall pray to non-obvious data leakage, and would benefit from more explainability. Some of its evaluation data overlap with the model’s own training inputs, raising the risk of circular benchmarking bias, and the study offers little insight into why the network makes its calls.

Leadership & Mentorship Reflection
Building trustworthy AI in medicine requires independent data, transparent reasoning, and humility about limitations that are baked into how these models work.
For early-career scientists, this paper is a case study in responsible ambition: innovate boldly, share your data openly, and interrogate your own benchmarks. AI or not, progress comes from rigorous, open science that understands its own limitations.