
This week’s paper is personal for me.
Dan Herschlag was my postdoc advisor. Dan taught me how to think mechanistically, how to test assumptions, and how to pursue scientific clarity with unflinching rigor.
That legacy is all over this paper. And is incredibly needed in this era of AI hype where data volume often sidelines careful model-driven science.
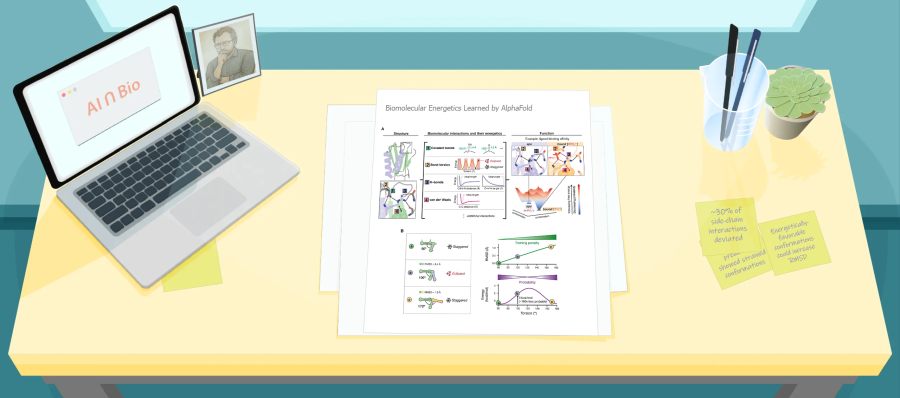
Herschlag et al. show that while AlphaFold3 predicts protein structures with backbone-level structural precision, it struggles to capture the physical rules that govern biological function.
But this paper isn’t a takedown: it’s a roadmap for how to go further.
What They Did
Rather than rely on RMSD alone, the team evaluated AlphaFold2 and AlphaFold3 against:
- Energetic rules: bond torsions, hydrogen bonds, and van der Waals contacts
- Experimental ensembles: from multi-temperature crystallography
- Model confidence: comparing pLDDT to physical plausibility
What They Found
- ~30% of side-chain interactions deviated from experimental observations, often with incorrect partners or implausible geometries
- High-confidence predictions (>90 pLDDT) still showed strained or physically invalid conformations
- Energetically-favorable conformations could increase RMSD and be penalized by the model
- AlphaFold3 missed ~85% of conformational variability seen in real experimental ensembles
Key Insight: Physics ≠ Proximity
AlphaFold can recapitulate Ramachandran and Lennard-Jones-like patterns. But that doesn’t mean it understands physical constraints.
To improve these tools, we need evaluation metrics grounded in molecular energetics, not just geometry.

Takeaway for Scientists
This paper is a reminder that progress requires more than prettier predictions. It demands models that reflect the physics that drive biology.